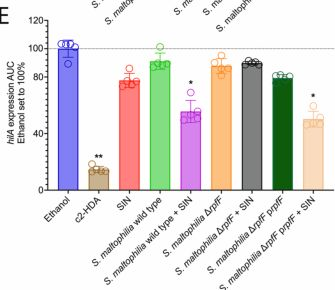
Successful colonization by enteric pathogens requires overcoming colonization resistance of the native microbiota while tightly regulating the expression of energetically-expensive virulence factors. Here we describe a feedback mechanism by which the enteric pathogen Salmonella orchestrates this balance through environmental manipulation. We show that Salmonella-induced oxidative stress can stimulate the colonic resident Stenotrophomonas maltophilia to enhance the secretion of the diffusible signal factor cis-2-hexadecenoic acid (c2-HDA), a potent repressor of Salmonella virulence. By sensing this metabolite, Salmonella can attenuate its own virulence program to favor proliferation and colonic colonization. In murine models, Salmonella colonization was significantly enhanced in the colon, and inflammation reduced, in the presence of c2-HDA produced by S. maltophilia. Moreover, the ability of Salmonella to recognize c2-HDA within the murine colon was crucial for its successful colonization. These findings reveal a pathogen-commensal signaling axis through which pathogen-driven inflammatory cues reshape the metabolic output of the microbiota, generating regulatory signals that are co-opted to optimize pathogen fitness in the gut.
Product Citations: 38

In Gut Microbes on 31 December 2026 by Chowdhury, R., Bosire, E. M., et al.
In Cell Reports on 27 January 2026 by Shirasaka, Y., Takeuchi, F., et al.
Viral protein genome-linked (VPg) is a small protein encoded by several non-enveloped positive-sense RNA viruses, including Cardiovirus. Although VPg is widely recognized as essential for viral RNA synthesis, its additional roles remain largely unexplored. Here, we report an alternative function of Cardiovirus VPg as an interferon-γ receptor agonist. During encephalomyocarditis virus (EMCV) infection, a member of the Cardiovirus genus, we uncovered that VPg from EMCV (VPgEMCV) is released extracellularly from host cells. VPgEMCV is then sensed by uninfected host cells, activating the interferon-γ signaling cascade and ultimately producing nitric oxide (NO) through inducible NO synthase induction. NO accumulation is crucial for triggering potent cell death in cooperation with tumor necrosis factor α (TNF-α) signaling, with TNF-α also being induced as an antiviral host response during EMCV infection. We demonstrate that VPgEMCV, comprising merely 20 amino acids, can exhibit cytokine-mimicry activity, with the synergistic interplay between VPgEMCV and TNF-α signaling regulating viral replication and disease pathogenesis.
Copyright © 2025 The Author(s). Published by Elsevier Inc. All rights reserved.
In Cancer Research on 2 September 2025 by Oleinik, N., Atilgan, F. C., et al.
Bioactive ceramide induces cell death in part by promoting mitophagy. C18-ceramide levels are commonly reduced in head and neck squamous cell carcinoma, which correlates with poor prognosis, suggesting the potential of harnessing ceramide for cancer treatment. In this study, we evaluated the ability of the ceramide analog D-erythro-14-(1-pyridinium)-N-octadecanoyl-sphingosine selenite (LCL768) to induce mitophagy and metabolic stress in head and neck squamous cell carcinoma. Mechanistically, LCL768 induced ceramide synthase 1 (CerS1)-mediated endogenous C18-ceramide accumulation in mitochondria to mediate mitophagy, which did not require the CerS1 transporter p17/PERMIT but was dependent on DRP1 activation via nitrosylation at C644. DRP1 facilitated the anchoring of the endoplasmic reticulum (ER) and mitochondrial membranes by promoting the association between phosphatidylethanolamine in the ER and cardiolipin in mitochondrial membranes. Mutations of Drp1 that prevented its binding to ER and mitochondrial membranes blocked CerS1/C18-ceramide mitochondrial accumulation, inhibiting LCL768-mediated mitophagy. In addition, LCL768-driven mitophagy altered mitochondrial metabolism, resulting in fumarate depletion and leading to tumor suppression in vivo. Exogenous fumarate supplementation prevented LCL768-mediated mitophagy, mitochondrial trafficking of CerS1, ER-mitochondrial tethering, and tumor suppression in mice. Fumarate metabolism was associated with PARKIN succination at a catalytic cysteine (Cys431), inhibiting its association with PINK1 and ubiquitin, thereby preventing mitophagy. LCL768-induced fumarate depletion attenuated PARKIN succination to promote PARKIN activation and mitophagy, indicating a feedforward mechanism that regulates mitophagy and fumarate metabolism through PARKIN succination. These data provide a mechanism whereby LCL768/CerS1-C18-ceramide-mediated mitophagy and tumor suppression are regulated by Drp1 nitrosylation, fumarate depletion, and PARKIN succination, providing a metabolic stress signature for lethal mitophagy.
The identification of a metabolic link between ceramide-induced mitophagy, fission, and fumarate depletion reveals an effective tumor suppressive strategy for head and neck squamous cell carcinoma.
©2025 American Association for Cancer Research.
In International Journal of Molecular Sciences on 25 July 2025 by Lu, Z., Zhou, X., et al.
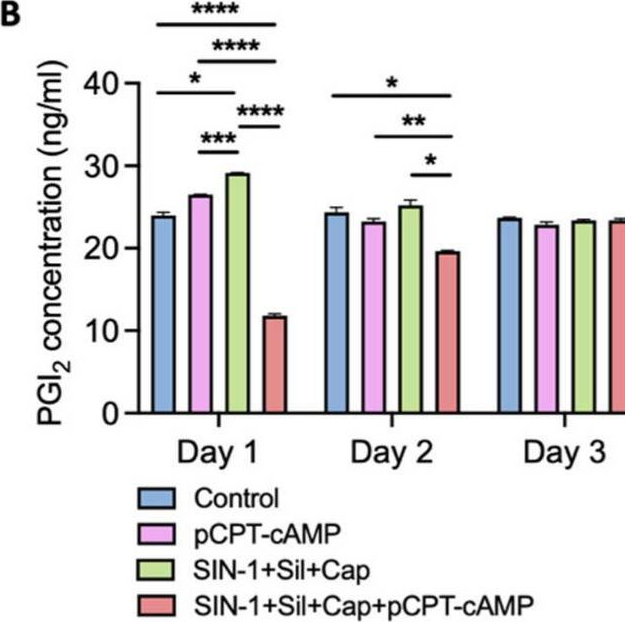
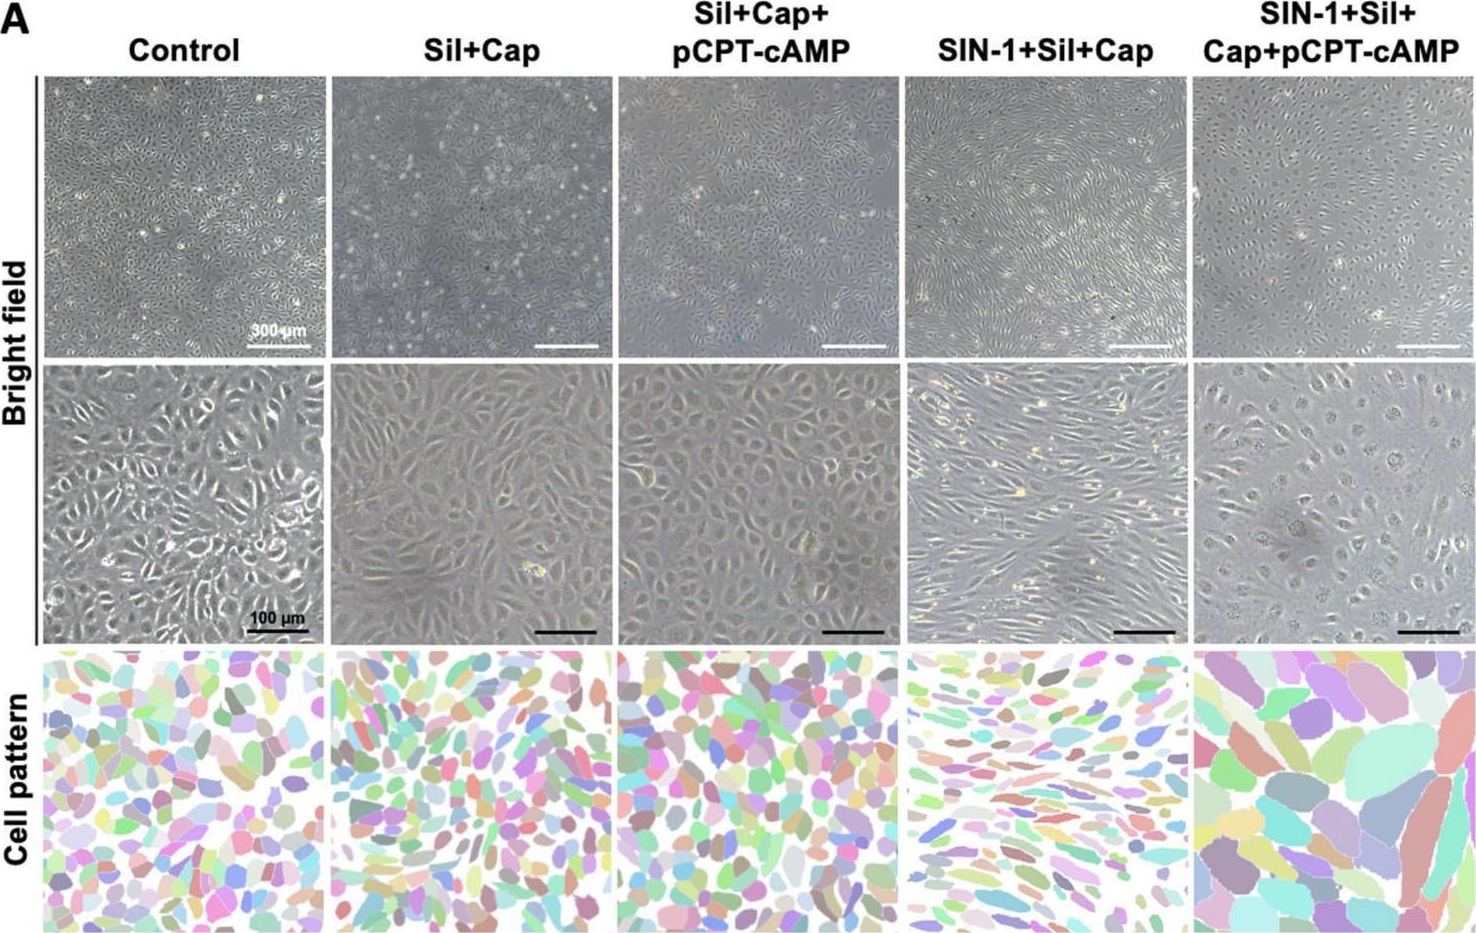
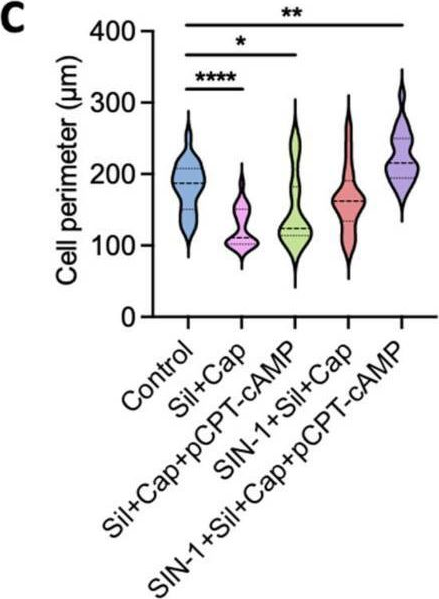
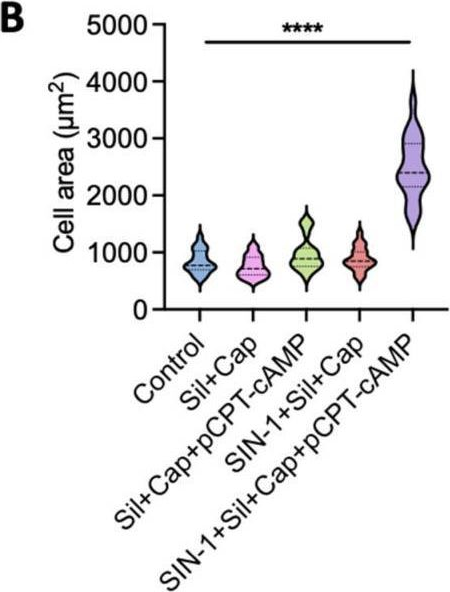
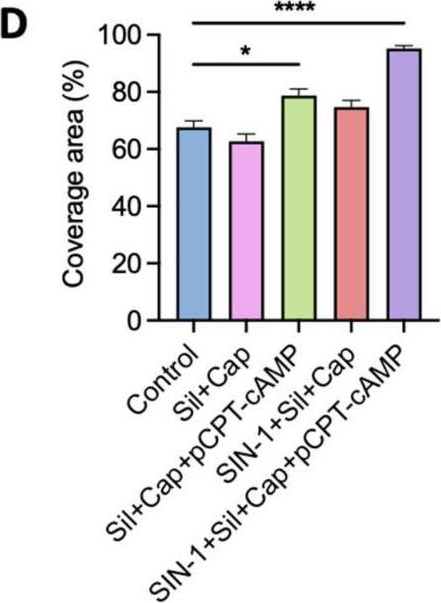
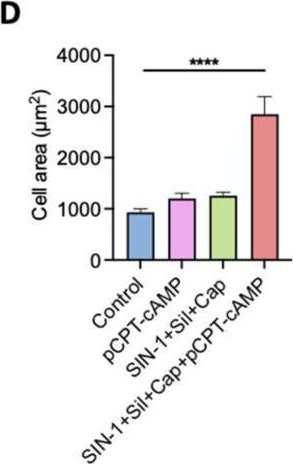
The development of functional endothelial monolayers on synthetic vascular grafts remains challenging, particularly for small-diameter vessels (<6 mm) prone to thrombosis. Here, we present a pharmacological strategy combining 8-(4-chlorophenylthio) adenosine 3',5'-cyclic monophosphate sodium salt (pCPT-cAMP, a tight junction promoter) with nitric oxide/cGMP pathway agonists 3-morpholinosydnonimine (SIN-1), captopril, and sildenafil) to enhance endothelialization. In human umbilical vein endothelial cells (HUVECs), this four-agent cocktail induced a flat, extended phenotype with a 3-fold increased cell area and 57.5% fewer cells required for surface coverage compared to controls. Immunofluorescence analysis revealed enhanced ZO-1 expression and continuous tight junction formation, while sustained nitric oxide (NO) production (3.9-fold increase) and restored prostacyclin (PGI2) secretion demonstrated preserved endothelial functionality. Anticoagulation assays confirmed a significant reduction in thrombus formation (p < 0.01) via dual inhibition of platelet activation and thrombin binding. These findings establish a synergistic drug combination that promotes rapid endothelialization while maintaining antithrombogenic activity, offering a promising solution for small-diameter vascular grafts. Further studies should validate long-term stability and translational potential in preclinical models.
Stable oxidative posttranslational modifications alter the gating properties of RyR1.
In The Journal of General Physiology on 2 December 2024 by Steinz, M. M., Beard, N. A., et al.
The ryanodine receptor type 1 (RyR1) is a Ca2+ release channel that regulates skeletal muscle contraction by controlling Ca2+ release from the sarcoplasmic reticulum (SR). Posttranslational modifications (PTMs) of RyR1, such as phosphorylation, S-nitrosylation, and carbonylation are known to increase RyR1 open probability (Po), contributing to SR Ca2+ leak and skeletal muscle dysfunction. PTMs on RyR1 have been linked to muscle dysfunction in diseases like breast cancer, rheumatoid arthritis, Duchenne muscle dystrophy, and aging. While reactive oxygen species (ROS) and oxidative stress induce PTMs, the impact of stable oxidative modifications like 3-nitrotyrosine (3-NT) and malondialdehyde adducts (MDA) on RyR1 gating remains unclear. Mass spectrometry and single-channel recordings were used to study how 3-NT and MDA modify RyR1 and affect Po. Both modifications increased Po in a dose-dependent manner, with mass spectrometry identifying 30 modified residues out of 5035 amino acids per RyR1 monomer. Key modifications were found in domains critical for protein interaction and channel activation, including Y808/3NT in SPRY1, Y1081/3NT and H1254/MDA in SPRY2&3, and Q2107/MDA and Y2128/3NT in JSol, near the binding site of FKBP12. Though these modifications did not directly overlap with FKBP12 binding residues, they promoted FKBP12 dissociation from RyR1. These findings provide detailed insights into how stable oxidative PTMs on RyR1 residues alter channel gating, advancing our understanding of RyR1-mediated Ca2+ release in conditions associated with oxidative stress and muscle weakness.
© 2024 Steinz et al.
In Gut Microbes on 31 December 2026 by Chowdhury, R., Bosire, E. M., et al.
Fig.3.E
Collected and cropped from Gut Microbes by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.5.B
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.3.A
-
IHC
-
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.3.C
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.3.B
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.3.D
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.4.D
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.5.E
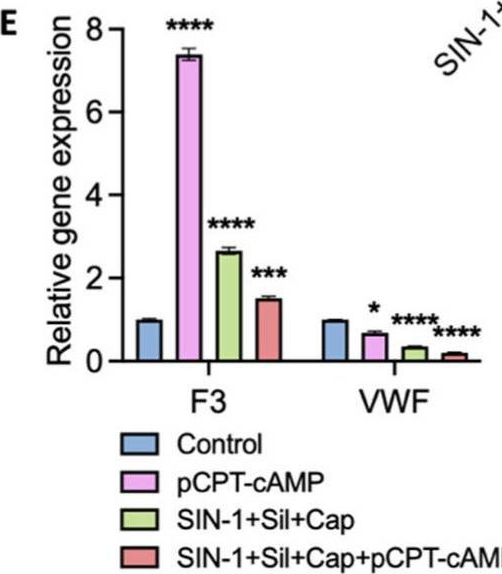
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.5.D
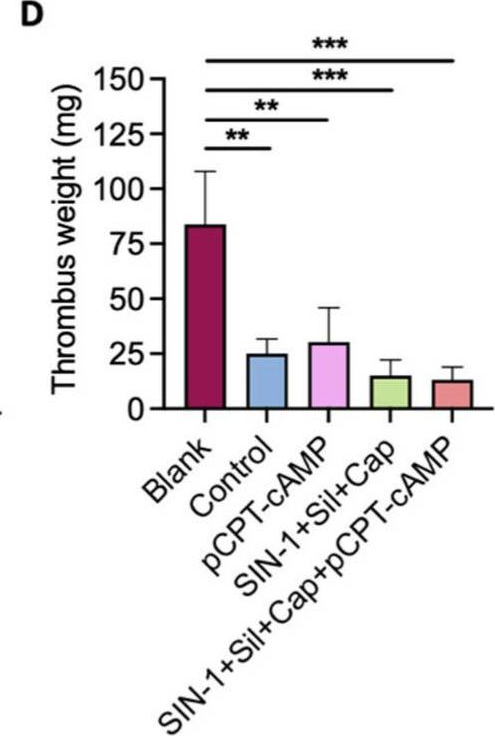
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.5.A
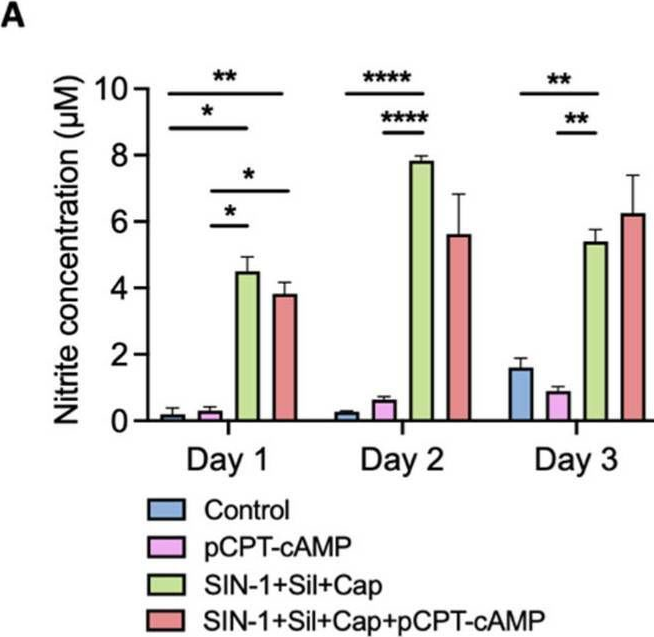
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.1.B
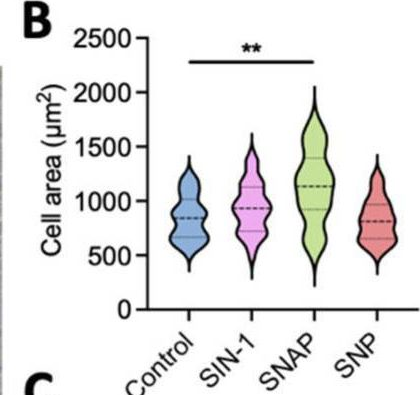
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.1.A
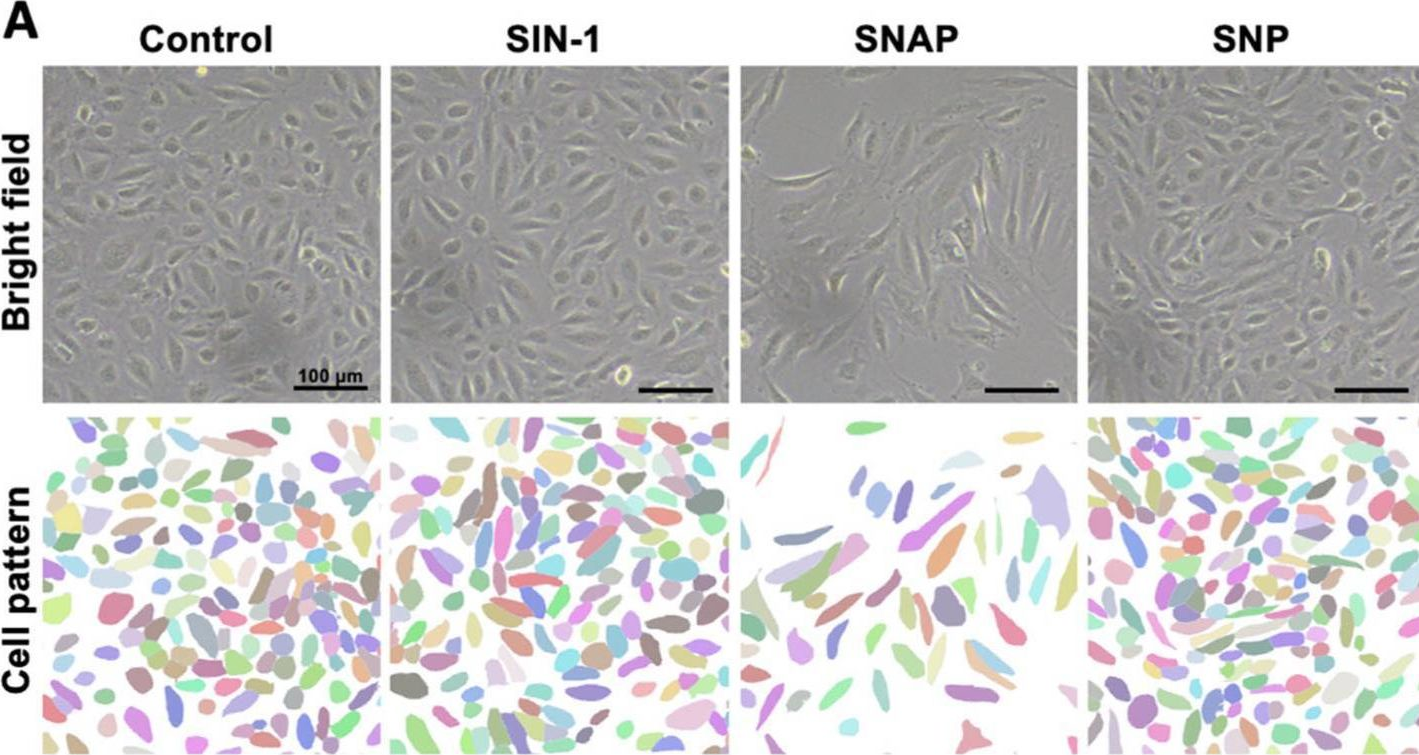
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.1.C
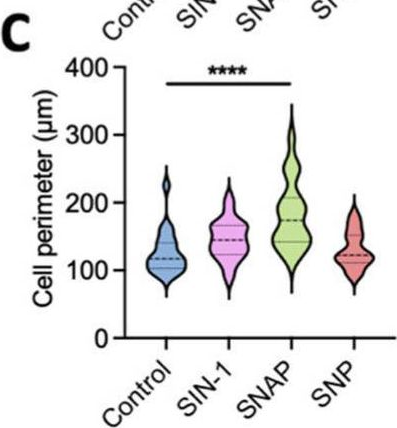
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.4.A
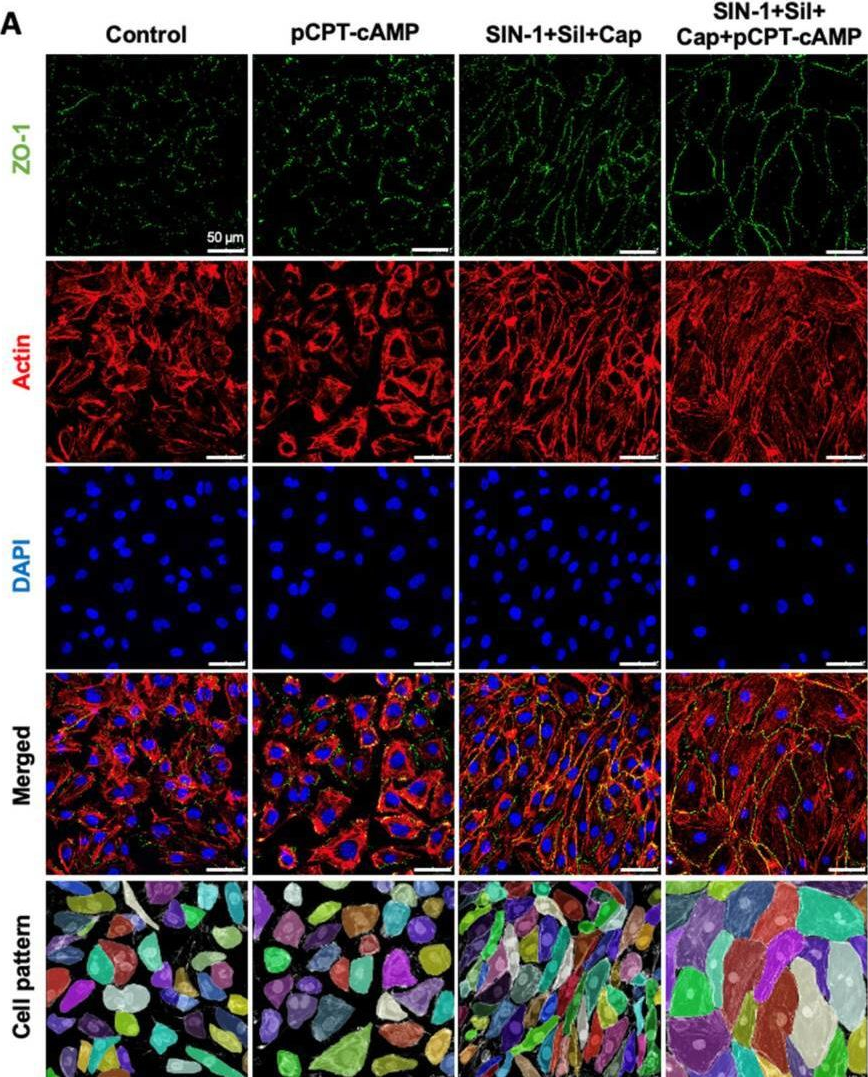
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.4.C
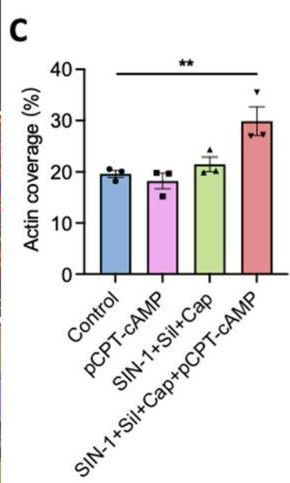
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.4.B
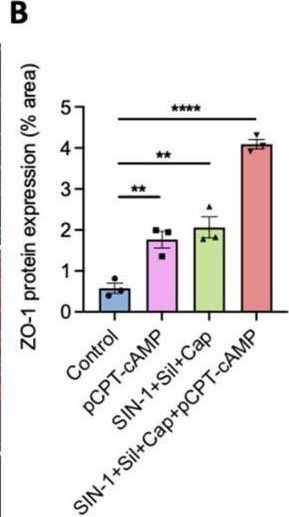
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.4.E
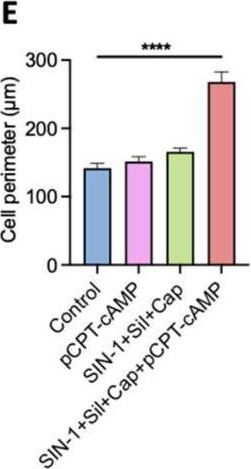
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 25 July 2025 by Lu, Z., Zhou, X., et al.
Fig.4.F
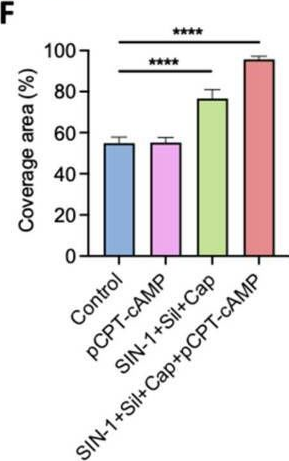
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In J Gen Physiol on 2 December 2024 by Steinz, M. M., Beard, N. A., et al.
Fig.1.A
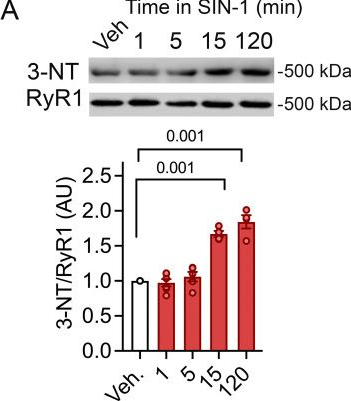
-
WB
-
Collected and cropped from The Journal of General Physiology by CiteAb, provided under a CC-BY license
Image 1 of 40
In J Gen Physiol on 2 December 2024 by Steinz, M. M., Beard, N. A., et al.
Fig.1.F
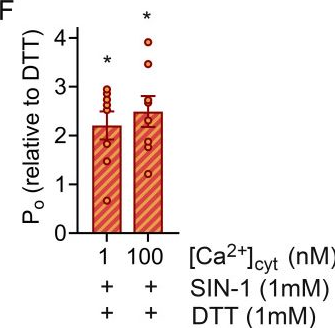
Collected and cropped from The Journal of General Physiology by CiteAb, provided under a CC-BY license
Image 1 of 40
In J Gen Physiol on 2 December 2024 by Steinz, M. M., Beard, N. A., et al.
Fig.1.E
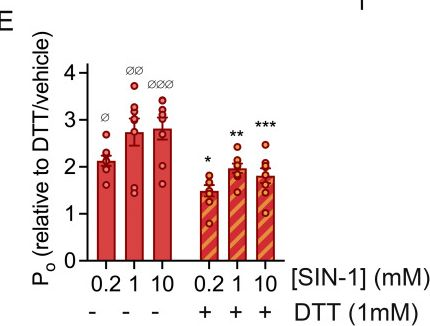
Collected and cropped from The Journal of General Physiology by CiteAb, provided under a CC-BY license
Image 1 of 40
In J Gen Physiol on 2 December 2024 by Steinz, M. M., Beard, N. A., et al.
Fig.1.D
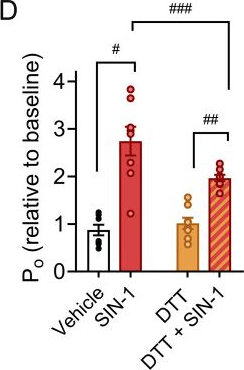
Collected and cropped from The Journal of General Physiology by CiteAb, provided under a CC-BY license
Image 1 of 40
In J Gen Physiol on 2 December 2024 by Steinz, M. M., Beard, N. A., et al.
Fig.3.D
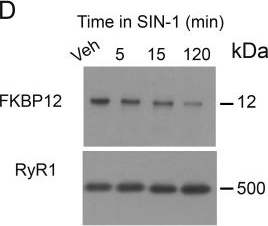
-
WB
-
Collected and cropped from The Journal of General Physiology by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 11 October 2021 by Lee, C. T., Hsieh, C. F., et al.
Fig.4.E
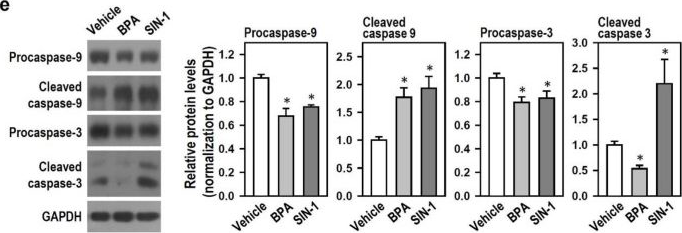
-
WB
-
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40
In Int J Mol Sci on 11 October 2021 by Lee, C. T., Hsieh, C. F., et al.
Fig.4.D
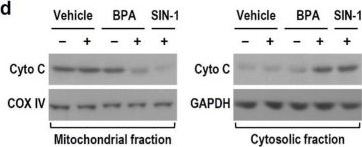
-
WB
-
Collected and cropped from International Journal of Molecular Sciences by CiteAb, provided under a CC-BY license
Image 1 of 40